Run TumorQuantAI
The beginner command-line interface expands a short command into the existing run.sh and Nextflow workflow while preserving the selected input, output, work directory, sampling, seed, source MPP, container, and model identity.
tumorquantai run
        |
        +-- inspect and validate inputs
        +-- select CPU, GPU, or prepared local profile
        +-- write a run manifest
        +-- start Nextflow
        +-- isolate every slide for retry and resume
        +-- aggregate completed slides and audit exclusions
Start with inspection
# Inspect the input roster without HistoPLUS inference.
tumorquantai inspect /path/to/slides \
  --output /path/to/tumorquantai-inspection \
  --source-mpp 0.261780
Open the inspection report and resolve missing L2 companions, duplicate warnings, unknown scale, or unexpected sample IDs before running.
Choose a preset
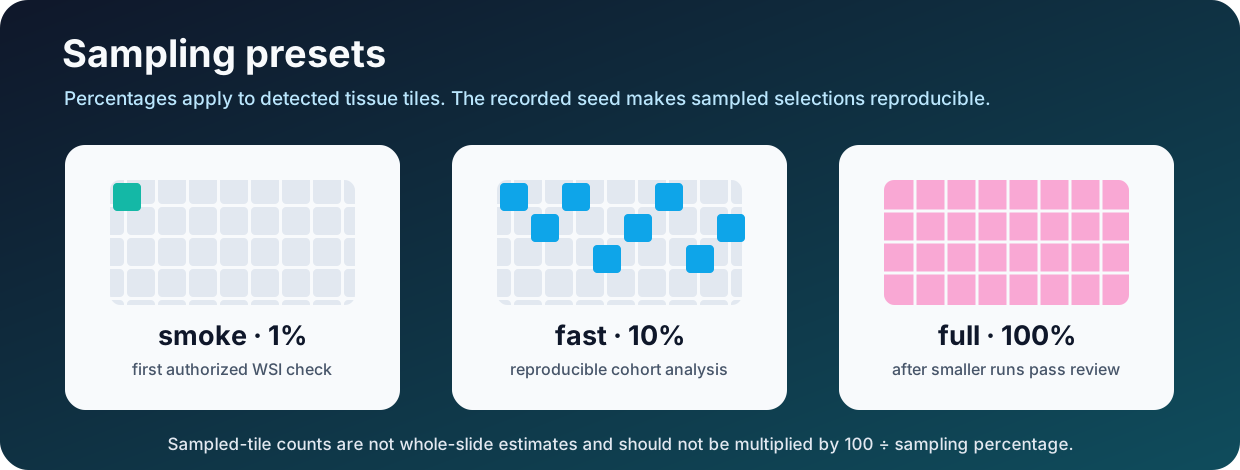
| Preset | Percent of detected tissue tiles | Typical use |
|---|---|---|
smoke |
1% from one selected slide | First authorized inference check |
fast |
10% | Reproducible exploratory cohort analysis |
full |
100% | Exhaustive processing after smaller runs pass review |
The random seed is recorded. The same input, preset, seed, and configuration select the same sampled tissue tiles.
Run one slide at 1%
# Run one selected sample on CPU.
tumorquantai run /path/to/slides \
  --sample-sheet /path/to/slides/samples.csv \
  --output /path/to/results-smoke \
  --work-dir /path/to/work-smoke \
  --preset smoke \
  --sample case_001 \
  --source-mpp 0.261780 \
  --cpu
Run a cohort at 10%
# Run all selected samples at a deterministic 10% on GPU.
tumorquantai run /path/to/slides \
  --sample-sheet /path/to/slides/samples.csv \
  --output /path/to/results-10-percent \
  --work-dir /path/to/work-10-percent \
  --preset fast \
  --source-mpp 0.261780 \
  --gpu
Use --cpu instead of --gpu when GPU execution is unavailable. CPU and GPU comparisons must use separate output and work directories.
Run all detected tissue
# Run every detected tissue tile only after smaller runs pass review.
tumorquantai run /path/to/slides \
  --sample-sheet /path/to/slides/samples.csv \
  --output /path/to/results-full \
  --work-dir /path/to/work-full \
  --preset full \
  --source-mpp 0.261780 \
  --gpu
full means 100% of detected tissue tiles, not every background pixel.
CPU, GPU, and local profiles
| Option | Behavior |
|---|---|
--cpu |
Maintained CPU container and CPU HistoPLUS execution |
--gpu |
Maintained GPU container; requires NVIDIA host and Docker runtime |
--profile auto |
Select GPU only when the host and Docker runtime are visible; otherwise CPU |
--profile local |
Expert-prepared local environment without Docker |
Run tumorquantai doctor before a new execution profile.
# Check a planned GPU result and work location.
tumorquantai doctor \
  --input /path/to/slides \
  --output /path/to/results-10-percent \
  --work-dir /path/to/work-10-percent \
  --online
Select samples
Use a sample sheet for exact sample identity. Add stable include or exclude globs when needed:
# Process cohort_A samples while excluding repeat samples.
tumorquantai run /path/to/slides \
  --sample-sheet /path/to/slides/samples.csv \
  --output /path/to/results-cohort-A \
  --work-dir /path/to/work-cohort-A \
  --preset fast \
  --source-mpp 0.261780 \
  --include 'cohort_A*' \
  --exclude '*repeat*' \
  --cpu
Use an explicit seed
# Run 10% with an explicit deterministic seed.
tumorquantai run /path/to/slides \
  --output /path/to/results-seed-20260709 \
  --work-dir /path/to/work-seed-20260709 \
  --preset fast \
  --source-mpp 0.261780 \
  --seed 20260709 \
  --cpu
Do not compare sampled runs that used different seeds without accounting for the changed tile selection.
Discovery-only dry run
# Validate discovery and print the expanded plan without inference.
tumorquantai run /path/to/slides \
  --output /path/to/results-dry-run \
  --preset fast \
  --source-mpp 0.261780 \
  --cpu \
  --dry-run
A dry run writes discovery and run-planning metadata but no biological prediction.
Stop and resume
Press Ctrl+C to stop. Repeat the exact command with the same output and work directories. Resume is enabled by default.
# Display current completion state and the exact local resume command.
tumorquantai status /path/to/results-10-percent
TumorQuantAI refuses to mix incompatible runs in one output directory. Use a new output root when changing:
- preset or percent;
- seed;
- source MPP;
- CPU/GPU/local profile;
- selected samples;
- expert passthrough arguments.
Do not move or delete an active Nextflow work directory. Do not run nextflow clean -f until outputs are verified and backed up.
Failure handling
Each slide runs independently. A slide can retry without rerunning completed slides. When continuation is enabled, other slides can finish after one sample fails.
A failed or incomplete sample:
- remains in
sample_aggregation_audit.csv; - has no numeric cohort-matrix column;
- is not interpreted as a biological zero;
- can be retried with the same command.
Generate the report
# Regenerate the portable run summary.
tumorquantai report /path/to/results-10-percent
Open START_HERE.html, then review overlays, summaries, the aggregation audit, and cohort tables.
Direct expert execution
The beginner CLI is recommended for routine use. The compatible expert interfaces remain available:
# Display the lower-level shell launcher options.
./run.sh --help

# Display the complete beginner and parameter-file options.
tumorquantai run --help
Direct nextflow run and run.sh usage require the user to manage protected parameters, model access, paths, resources, and provenance correctly. See CLI reference, Parameters, and Configuration.